In an article at the Nautilus web site, scientist Ken Richardson suggests that his fellow scientists have been guilty of some molecular mythology. He points out that scientists have repeatedly used “action verbs” in describing DNA, telling us that inside DNA are genes that “act,” “behave,” “direct,” “control,” “design,” are “responsible for,” and so forth. But then Richardson tells us “a counter-narrative is building” to correct such erroneous ideas, and then gives us reasons for thinking that genes are merely passive chemical units that do no such things.
Another example of molecule mythology involves a protein called PKMzeta. Some neuroscientists have suggested that PKMzeta has the ability to make memories last for decades in synapses, even though the proteins that make up synapses are very short-lived (having an average lifetime of two weeks or less). Quite a few of the papers or posts spreading this idea were written or co-written by the same person, Todd C. Sacktor. It is never explained clearly by any such theorists how a protein molecule could perform this great feat of magic. For anyone to explain such a thing clearly, he would first need to have a clear theory of how conceptual memories and episodic memories could be stored in synapses. No neuroscientist has ever presented a clear and explicit theory of any such thing. Neuroscientists merely vaguely tell us that somehow memory storage in a brain occurs through “synapse strengthening,” without presenting any clear theory of how that could be memory storage.
Of course, if you do not have a clear theory of how memories could be stored (for even a few minutes) in synapses, you cannot possibly have a clear theory as to how some protein molecule such as PKMzeta could possibly cause memories stored in synapses to persist for decades, even though the proteins that make up such synapses are very short-lived, lasting an average of less than two weeks. Trying to defend against the charge that synapses are totally unsuitable for storing memories for decades, because of the short lifetimes of the proteins that make up synapses, a scientific paper states, “As long as PKMZ [PKMzeta] remains active and there is an absence of forces which terminate its activity (such as LTD), it will continue to sustain the biochemical changes at the synapse which serve as the neurobiological basis of memory, allowing the memory to persist for durations far exceeding the turnover of its component molecules.” But how could such a miracle of persistence occur, which would be like a message written in wet sand at the seashore persisting for decades, even though the wet sand was being replaced and written over whenever the tide came in? The science paper does not tell us.
Again, we have the case of an “action verb” inappropriately used to describe a molecule. We are told that PKMzeta has a “sustain” super-power allowing it to preserve fantastically complicated information supposedly stored externally in synapses made up of short-lived molecules that are constantly being replaced. There is nothing in the structure of PKMzeta that should cause us to believe it can do any such thing. No theorist has presented an explicit theory as to how anything like PKMzeta could preserve a memory. Such theorists may sometimes present chemical details to impress us, but such details do not constitute a theory unless a theorist gives explicit examples of precisely how specific memories (such as someone's memory of seeing Paris or someone's memory of details learned about World War I) could be permanently stored with the aid of PKMzeta. No theorist has done any such thing.
I suppose that if a PKMzeta molecule were able to cause memories to persist despite rapid protein turnover, we might imagine it as some kind of "genius" molecule that has thoughts like this:
Oh, my goodness, I see that a memory is starting to degrade because of protein turnover! The memory now states, "Ottawa is the capitol of," which isn't even a full English sentence. Why, I'd better synthesize some new proteins to fill in for those proteins that died, so there can be a nice complete English sentence. Now, what was that country that Ottawa is the capital of?
Of course, anything the slightest bit like this is very hard to believe in. It would seem that the most minimal requirements that a molecule would have to fulfill in order to be a "memory maintenance molecule" would be the following:
(1) The molecule would have to somehow know whenever a particular protein molecule (that was part of a memory stored in a synapse) had died or disappeared because of the short lifespans of protein molecules.
(2) The molecule would have to somehow cause a replacement protein of the same type to appear in the same place as the vanished molecule, so that the memory did not degrade.
The problem is that no one can envision a credible scenario under which a molecule could have either of these powers. To imagine how much of a miracle it would be for memories to persist despite constant protein turnover, you can imagine a homeowner with ten picnic tables in his backyard, each of which is filled with leaves on which a word or two is written. Imagine these leaves spell out narratives, factual information, and ideas. But the problem is that about one day in three there are winds blowing the leaves off of the tables, and scattering them far away. Also, the leaves don't last longer than a year, because they tend to crumble. Now imagine the homeowner has to keep all this information preserved in the leaves, not just for a few nights but for 50 years. That would be a mountainous job. An equally mountainous job would have to be done if memories were to be preserved in brains despite constant protein turnover causing proteins to persist an average of less than two weeks, and no one has explained how a molecule could possibly do such a feat. Since synapses face not only rapid protein turnover inside them but also the problem that synapses don't last for longer than a year or two, they have the same "double degradation" problem that such a homeowner would have with his information written on leaves.

In the article here, a PKMzeta enthusiast is asked to explain how PKMzeta could cause memories to persist. The scientist gives a lengthy answer which fails to explain how PKMzeta could do such a thing. He merely says "a cluster of PKMzeta molecules can keep themselves turned on perpetually," and then claims that this supposed ability "is a plausible mechanism for memory persistence," without justifying that claim. This fragmentary theorizing is just hand waving. It has never been demonstrated that any cluster of PKMzeta molecules is capable of storing any information (such as a list of words) for a period as long as a month. We can imagine hypothetical lab experiments that might try to show such a thing, but they have never been done. The paper here refers to "900 synaptic proteins." PKMzeta is only one of those 900 proteins in synapses, being no more common in synapses than an average synapse protein. You don't solve the "short lifetime of proteins" problem by trying to argue that one in 900 of those proteins might somehow have some stability. As for the scientist's use of the word "plausible," it has been noted by others that "plausible" is the most abused word in theoretical science discourse, and that scientists often carelessly use the word "plausible" without ever doing anything at all to show a likelihood.
I suppose that if a PKMzeta molecule were able to cause memories to persist despite rapid protein turnover, we might imagine it as some kind of "genius" molecule that has thoughts like this:
Oh, my goodness, I see that a memory is starting to degrade because of protein turnover! The memory now states, "Ottawa is the capitol of," which isn't even a full English sentence. Why, I'd better synthesize some new proteins to fill in for those proteins that died, so there can be a nice complete English sentence. Now, what was that country that Ottawa is the capital of?
Of course, anything the slightest bit like this is very hard to believe in. It would seem that the most minimal requirements that a molecule would have to fulfill in order to be a "memory maintenance molecule" would be the following:
(1) The molecule would have to somehow know whenever a particular protein molecule (that was part of a memory stored in a synapse) had died or disappeared because of the short lifespans of protein molecules.
(2) The molecule would have to somehow cause a replacement protein of the same type to appear in the same place as the vanished molecule, so that the memory did not degrade.
The problem is that no one can envision a credible scenario under which a molecule could have either of these powers. To imagine how much of a miracle it would be for memories to persist despite constant protein turnover, you can imagine a homeowner with ten picnic tables in his backyard, each of which is filled with leaves on which a word or two is written. Imagine these leaves spell out narratives, factual information, and ideas. But the problem is that about one day in three there are winds blowing the leaves off of the tables, and scattering them far away. Also, the leaves don't last longer than a year, because they tend to crumble. Now imagine the homeowner has to keep all this information preserved in the leaves, not just for a few nights but for 50 years. That would be a mountainous job. An equally mountainous job would have to be done if memories were to be preserved in brains despite constant protein turnover causing proteins to persist an average of less than two weeks, and no one has explained how a molecule could possibly do such a feat. Since synapses face not only rapid protein turnover inside them but also the problem that synapses don't last for longer than a year or two, they have the same "double degradation" problem that such a homeowner would have with his information written on leaves.

In the article here, a PKMzeta enthusiast is asked to explain how PKMzeta could cause memories to persist. The scientist gives a lengthy answer which fails to explain how PKMzeta could do such a thing. He merely says "a cluster of PKMzeta molecules can keep themselves turned on perpetually," and then claims that this supposed ability "is a plausible mechanism for memory persistence," without justifying that claim. This fragmentary theorizing is just hand waving. It has never been demonstrated that any cluster of PKMzeta molecules is capable of storing any information (such as a list of words) for a period as long as a month. We can imagine hypothetical lab experiments that might try to show such a thing, but they have never been done. The paper here refers to "900 synaptic proteins." PKMzeta is only one of those 900 proteins in synapses, being no more common in synapses than an average synapse protein. You don't solve the "short lifetime of proteins" problem by trying to argue that one in 900 of those proteins might somehow have some stability. As for the scientist's use of the word "plausible," it has been noted by others that "plausible" is the most abused word in theoretical science discourse, and that scientists often carelessly use the word "plausible" without ever doing anything at all to show a likelihood.
But the PKMzeta enthusiasts have done a few studies which they claim lends credibility to their claims. I will describe a typical such study. A small number of mice are injected with something that suppresses the PKMzeta molecules in their body (or perhaps they are genetically engineered so that they don't have any PKMzeta). Memory experiments are then done. It is sometimes claimed that such mice perform not as well as normal mice. Such experiments have been hailed as support for the “memory maintenance” claims about PKMzeta.
There are several reasons why such studies do not at all show the claims about PKMzeta are correct. The first is that a result such as I described could never show that PKMzeta can save memories from destruction for years. Whenever memory is tested, it's hard to figure out what the cause is for a discrepancy between two test groups. A difference in a test result might be because (1) PKMzeta is involved in perceiving whatever observation is being tested; (2) or that PKMzeta is involved in memory storage; (3) or that PKMzeta is involved in memory retrieval; (4) or that PKMzeta has something to do with attention or focus or visual perception used in a memory test. A test discrepancy could never tell us which of these things was involved. And if some mice did worse in remembering things without PKMzeta, that might justify the small claim that PKMzeta has something to do with memory, but could never justify the vastly more extravagant claim that PKMzeta is capable of preserving memories for decades.
Another reason why such studies do not at all show the claims about PKMzeta are correct has to do with a general malaise in neuroscience. A general problem in modern neuroscience is the production of papers with marginal results that we cannot trust because of things such as too-small sample sizes and publication bias. Let us imagine that neuroscientists want to prove some idea that fits in with their ideological expectations. A great number of experiments might be done, almost all producing no support for the idea. But perhaps 1 in 20 might produce results marginally supporting the idea, probably because of chance variations in data. Now, today negative results are vastly less likely to get published than positive results. So if 19 researchers get a negative result, conflicting with what neuroscientists hope to get, it could be that 10 of them don't even bother to write up their results as a scientific paper, and that the other 9 do write up a paper but don't get it published (because of the journal bias against negative results). However the one researcher who (by chance) got a positive result will write up his result as a scientific paper. Since it will be a result neuroscientists were hoping to get, he will almost certainly get the result published.
This publication bias is a great problem affecting the reliability of scientific research. Because of it we should follow a precautionary neuroscience rule like this: don't believe something has been established unless the result turns up fairly consistently at a high level of significance, in studies with large sample sizes.
Has this happened in regard to memory experiments involving PKMzeta? Not at all. In 2011 a scientist reported three separate studies showing that inhibiting PKMzeta has no effect on memory if tested between 10 and 15 day after the memory forms. In 2013 two groups of scientists published results conflicting with claims that PKMzeta might allow memories to persist a long time. One study by a team of scientists used genetically engineered mice that had no PKMzeta. It found that such mice “have no deficits in several hippocampal-dependent learning and memory tasks,” and concluded that PKMzeta is not required for memory or learning. Another study by a different team of scientists found that absence of PKMzeta “does not impair learning and memory in mice.” A 2015 study found that inhibiting PKMzeta has no effect on memory in tests performed 30 days after the memory forms. A 2016 paper also found that that inhibiting PKMzeta has no effect on memory in tests performed 30 days after the memory forms.
Such studies would seem to completely debunk claims that PKMzeta enables memories to persist for decades in synapses despite the short lifetimes in the proteins.
The scientists such as Sacktor who helped to spread the PKMzeta myth have tried to fight back with papers such as this 2016 paper. But in that very paper we see evidence that second-rate science is being used to try to prop up claims about PKMzeta. In Figure 7 the scientists tell us how many mice were used for their experiment involving the memory effects of PKMzeta deprivation. They used only 8 mice per study group. That's way too small a sample size to get a moderately convincing result. It is well known that at least 15 animals per study group should be used to get a moderately convincing result. If you use only 8 animals per study group, there's a very high chance you'll get a false alarm, in which the result is due merely to chance variations rather than a real effect in nature. In fact, in her post "Why Most Published Neuroscience Studies Are False," neuroscientist Kelly Zalocusky suggests that neuroscientists really should be using 31 animals per study group to get a not-very-strong statistical power of .5, and 60 animals per study group to get a fairly strong statistical power of .8. Compare these numbers to the 8 animals per study group mentioned in Figure 7 of the Sacktor paper.
This is the same “too small sample size” problem (discussed here) that plagues very many or most neuroscience experiments involving animals. Neuroscientists have known about this problem for many years, but year after year they continue in their errant ways, foisting upon the public too-small-sample-size studies with low statistical power that don't prove anything because of a high chance of false alarms.
If you look up the PRKCZ gene behind the PKMZeta protein molecule, using this page and this page of the Human Protein Database, you will find no characteristics that seem unusual, and nothing suggesting any superstar status. The pages make no mention of the gene even being used in synapses, telling us that the gene is "mainly localized to the cytosol" and "in addition localized to the plasma membrane." The very idea of some kind of "superstar protein" or "superstar gene" is contrary to the experience in recent decades of scientists, who have found in general that bodily functions almost always involve the coordinated ballet of very many different genes (typically hundreds of them to accomplish a particular task).
If you look up the PRKCZ gene behind the PKMZeta protein molecule, using this page and this page of the Human Protein Database, you will find no characteristics that seem unusual, and nothing suggesting any superstar status. The pages make no mention of the gene even being used in synapses, telling us that the gene is "mainly localized to the cytosol" and "in addition localized to the plasma membrane." The very idea of some kind of "superstar protein" or "superstar gene" is contrary to the experience in recent decades of scientists, who have found in general that bodily functions almost always involve the coordinated ballet of very many different genes (typically hundreds of them to accomplish a particular task).
The 2015 scientific paper here shows that PKMzeta rapidly degrades in synapses. The authors say that therefore a stable amount of PKMzeta "would be difficult to maintain at synapses and store memories over long time scales." The paper tells us “There is growing evidence against a role for PKMzeta in memory.” Figure 9 of the paper also shows that a kind of cousin molecule or "isoform" of PKMzeta (PKC lambda) also quickly degrades, experiencing a 50% loss or degradation every 10 hours. So it seems that there is no truth to the idea of PKMzeta (or PKC lambda) as some magic bullet that allows memories to persist for decades in synapses that are constantly having their proteins replaced.
Where does that leave neuroscientists? It leaves them without a leg to stand on in their claims that memories are stored in brains. Based on everything we know about synapses, there is no reason to believe that synapses are capable of storing a memory for even a month, let alone the 50 years that is how long older humans can remember things. As discussed here and here, equally grave problems prevent scientists from creating any credible account of how memories could be encoded into neural states or how seldom-retrieved facts learned many years ago could be instantaneously recalled from a brain that seems to lack any capability for fast look-ups from exact neural positions. We also know (as discussed here and here) that massive damage can occur to brains (such as surgical removal of half of a brain) while producing little effect on memory, which would seem to be impossible if memories are stored in brains. How long before we realize that human memory cannot be a neural thing, but must be a psychic or spiritual phenomenon?
Some people tell tall tales about the protein CAMKII similar to the tall tales told about PKMZeta. We are sometimes told that some alleged autophosphorlyation of CAMKII can help explain stable memories. Most of the reasons I have cited against PKMZeta also apply with equal strength to CAMKII. At this link we are told an experiment debunked the idea that autophosphorlyation of CAMKII has a role in memory storage. The lifetime of a CAMKII molecule is only 30 hours, according to this source. The book here makes this statement:
Some people tell tall tales about the protein CAMKII similar to the tall tales told about PKMZeta. We are sometimes told that some alleged autophosphorlyation of CAMKII can help explain stable memories. Most of the reasons I have cited against PKMZeta also apply with equal strength to CAMKII. At this link we are told an experiment debunked the idea that autophosphorlyation of CAMKII has a role in memory storage. The lifetime of a CAMKII molecule is only 30 hours, according to this source. The book here makes this statement:
"In the mid-1980's there was much excitement about the idea that autophosphorlyated CaMKII might serve as a self-perpetuating signal that could subserve permanent memory storage. However, a variety of experimental results generated since then suggests that perpetual activation of CaMKII does not occur with LTP-inducing stimulation or memory storage."
This scientific paper says the following:
"Previous models have suggested that CaMKII functions as a bistable switch that could be the molecular correlate of long-term memory, but experiments have failed to validate these predictions....The CaMKII model system is never bistable at resting calcium concentrations, which suggests that CaMKII activity does not function as the biochemical switch underlying long-term memory."
This recent scientific paper says on page 9, "Overall, the studies reviewed here argue against, but do not completely rule out, a role for persistently self-sustaining CaMKII activity in maintaining" long term memory. Another paper says, "The autophosphorylation of CaMKII, once thought to help maintain long-term changes in synaptic strength, has since been revealed to be rather transient."
This recent scientific paper says on page 9, "Overall, the studies reviewed here argue against, but do not completely rule out, a role for persistently self-sustaining CaMKII activity in maintaining" long term memory. Another paper says, "The autophosphorylation of CaMKII, once thought to help maintain long-term changes in synaptic strength, has since been revealed to be rather transient."
Those who have studied the history of science are familiar with epicycles, a complicated speculation that was introduced into Ptolemy's theory of astronomy, to try to fix cases in which the theory did not match observations. We may say these CaMKII speculations and PKMZeta speculations are epicycles intended to fix the failing synaptic theory of memory storage. But while the Ptolemaic epicycles were exact speculations, the CaMKII speculations and PKMZeta speculations are very vague, failing to specify any exact theory of memory storage.
Last week a press release from New York University tried to breath life into the dead horse of the mythology of "memory maintenance molecules." The press release was a glaring example of what constantly occurs these days in university press releases: university PR offices boasting about grand accomplishments that were not actually accomplished. We have the untrue claim that a "new study in the journal Science Advances, conducted by a team of international researchers, has uncovered a biological explanation for long-term memories." No, the study was just more Questionable Research Practices junk science, a study so poorly designed it is a wonder it got published. We read this:
"It’s been long-established that neurons store information in memory as the pattern of strong synapses and weak synapses, which determines the connectivity and function of neural networks. However, the molecules in synapses are unstable, continually moving around in the neurons, and wearing out and being replaced in hours to days, thereby raising the question: How, then, can memories be stable for years to decades?"
No, it has not ever been established that " neurons store information in memory as the pattern of strong synapses and weak synapses" : no credible theory of how such a thing could work has ever been advanced, no memory information stored in neurons or synapses has ever been found through microscopic examination, and the instability of synapses (including the short lifetimes of their proteins) is the strongest reason for thinking that it cannot possibly be true that "neurons store information in memory as the pattern of strong synapses and weak synapses."
The press release makes the groundless claim that the junk science paper it is publicizing shows "that KIBRA is the 'missing link' in long-term memories," referring to a molecule called KIBRA. The press release makes the groundless claim that "their experiments in the Science Advances paper show that breaking the KIBRA-PKMzeta bond erases old memory."
The study is the study "KIBRA anchoring the action of PKMζ maintains the persistence of memory" which you can read here. We have a very bad example of Questionable Research Practices junk science. The study group sizes used are ridiculously small, with study groups as small as only 4 mice and 6 mice, and the average study group size being only about 7 mice. It is frequently pointed out to neuroscientists that experimental studies involving mice are generally worthless unless they use at least 15 subjects per study group; but neuroscientists keep senselessly continuing to use ridiculously low study group sizes. Why do they do that? Because it allows them to "mine noise," and report false alarms that would vanish if a decent study group size was used. It's rather like someone trying to prove his prophetic powers by publishing a test in which he correctly predicted whether merely four consecutive coin flips were "heads" or "tails," conveniently failing to publish a larger test of his powers involving how well he predicted 15 consecutive coin flips. You can get all kinds of false alarms when you use tiny sample sizes.


Attempting to measure whether a rodent remembered something fearful by doing estimates of "freezing behavior" is not a reliable way of measuring fear or memory, but instead an unreliable "see whatever you want to see" affair. All studies hinging on so unreliable a method are junk science studies, including the new study by Sacktor mentioned in the New York University press release. Contrary to the claims by Sacktor in his later paper and the claims in the New York University press release, no robust evidence has been produced to show that inhibiting either the KIPRA molecule or the PKMζ molecule (PKMzeta) does anything to harm the memory of mice. In the paper and the press release Todd Sacktor tells the tall tales of memory maintenance mythology that he likes to tell, unbelievable tales that are not backed up by any robust experiments.
Genuine "freezing behavior" in an animal would typically only be an instantaneous thing, lasting only a few seconds. The longer the length of time over which "freezing behavior" is judged, the more unreliable such a judgment is as a basis for judging whether the animal recalled a fearful stimulus. In Sacktor's latest paper discussed above, which you can read here, fear recall is being judged by someone estimating how much an animal was non-moving over a length of four minutes. That's a particularly unreliable use of the utterly unreliable technique of judging "freezing behavior" as a method of trying to determine whether recall occurred.
A 2024 scientific paper makes this candid confession, using the phrase "still not completely understood" when it should be saying "are not at all understood":
"Despite over a hundred years of research, the cellular/molecular mechanisms underlying learning and memory are still not completely understood. Many hypotheses have been proposed, but there is no consensus for any of these."
The paper calls Sacktor's speculations about the PKMζ molecule (PKMzeta) "very controversial."
I can give you an example that helps show the difficulty of maintaining stable information from unstable components. Let's suppose a shaving creme company wants to publicize its product. It arranges for a line of people to appear outside of the main branch of the New York Public Library, a line in which each person will be displaying a letter made out of shaving creme. The total line of people will spell out the message: "Sale! 10% off on purchases of Barbisol shaving creme, the world's most comfortable shaving creme." The letters will look like this, but each letter will rest on the outstretched palms of one person.

(1) A system for representing words by use of symbolic tokens (the English alphabet).
(2) Some skill for creating these tokens in shaving creme letters (maybe someone who has practiced this skill).
(3) An addressing system by which each person in the line knows his ordinal position in the line.
(4) A message system by which components that are about to fail send out a message to some receiving system notifying it to replace their failing token, telling that receiving system of which token to replace, and what the address was of the token to replace.
No similar system could ever be possible in the human brain. The human brain has no addresses or position numbers or position notation system or coordinate system. Neurons and synapses have no knowledge of the English alphabet, and no capability of writing synapse states or neuron states corresponding to letters of the English alphabet. A protein about to decay in a synapse would never know it was about to decay, and would never be capable of sending some external receiver a "replace me" message. And such a message could never have the address or position coordinates of a synapse protein to be replaced, because tiny components in the brain have neither addresses nor position coordinates. When you also consider that synapses are all entangled in 3D space, making them geometrically more difficult to work with than a simple one-dimensional line, you may start to realize how mythical is the notion that stable memories lasting decades could be made from constantly-replaced components with lifetimes of only a few weeks.
Postscript: A 2018 paper tells us a little about the appalling state of research practices in research involving rodents:
"There is a crisis in pre-clinical biomedical research
involving laboratory animals. Too many papers publish
results which turn out to be irreproducible. One estimate puts the cost at $28 billion being wasted per annum in the United States alone. The causes of this irreproducibility crisis have not
been fully identified. But it has been known for many
years that experiments are often poorly designed, inadequately analysed, and misreported. A survey of
271 papers chosen at random involving rats, mice
and non-human primates showed that 87% did not
report random allocation of experimental subjects to
the treatments and 86% did not report 'blinding'
when measuring the results. None of the papers gave
any justification for their choice of sample size, and a
substantial number of papers failed even to state the
sex, age or weight of the animal."
A 2021 paper ("Increasing the statistical power of animal experiments with historical control data" by V. Bonapersona et. al. ) gives us the damning graph below:
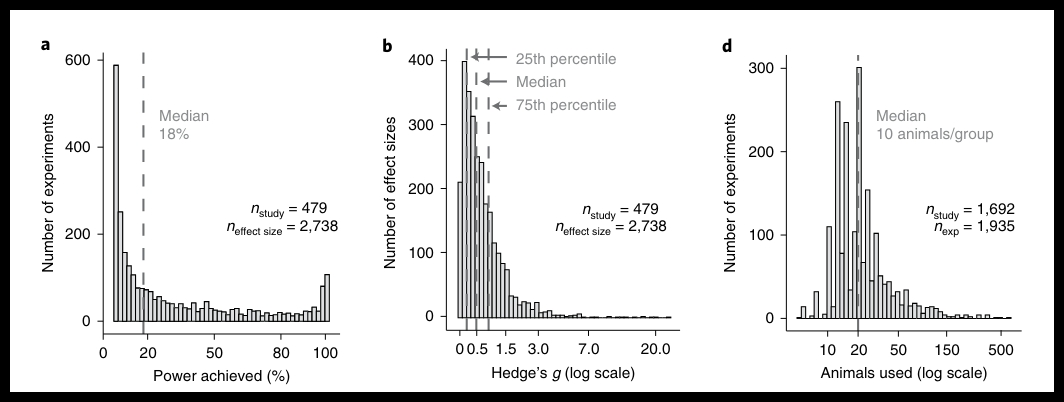
"For a common effect size of Hedge’s g= 0.5
(Welch’s independent samples t-test, α=0.05), ten animals per
group would correspond to a statistical power of 18%, 30 animals
per group to 48% power and 65 animals per group to 81% power...Through a systematic search (Supplementary Notes 1 and 2), we identified a large sample of animal studies in the areas of ‘neuroscience’ and ‘metabolism’ (n...=1,935) that were previously included in meta-analyses (n...=69). These animal studies had an overall median statistical power of 18% (Fig. 1a), which was roughly equal in the two fields (neuroscience, 15%; metabolism, 22%)....We estimated that, at best, 12.5% of a large sample of rodent studies were sufficiently powered (that is, prospective power was larger
than 80%). This estimate is a best-case scenario, as it is not yet
adjusted for any subsequent multiple testing, experimental bias, P
hacking and/or fishing, selective reporting, etc.."
Postscript: Scientific American has an article covering Sacktor's latest paper, giving us yet another example of its very poor journalism regarding neuroscience research. We have an appalling failure to inform the reader of the most basic facts about Sacktor's latest study. At no point is the reader told that the research merely involved rodents. At no point are we told about the appallingly small study group sizes used, such as only 7 rodents. Sacktor is allowed to get away with a groundless boast that he "nailed it," and no mention is made of how the study hinged upon an unreliable technique for measuring memory (the faulty judgement of "freezing behavior" discussed above). The author falls for Sacktor's boasts "hook, line and sinker."
I may also mention that the study fails to discuss how an effective blinding protocol was followed. We get a mere sketchy vague statement that an experimenter judging how much "freezing behavior" occurred was "blind to the conditions," but that does not constitute a description of an effective blinding protocol. When very small study group sizes (such as only seven mice) are used it is easy for a supposedly blind judge to know (by visual recognition) whether or not some animals being analyzed were part of some group that had been chemically treated, and which were desired to be described as acting in a particular way, such as "freezing" more. Effective blinding in an experiment (necessary whenever subjective judgments are made) can only be achieved by following a careful, detailed blinding protocol taking at least a long paragraph to describe; and that apparently did not occur in this study. No reliable measurement or reliable analysis of memory performance has occurred.
Post-postscript: A perennial promoter of junk neuroscience research, Quanta Magazine had a 2025 article promoting Sacktor's groundless boasts, one with the bogus title "The Molecular Bond That Helps Secure Your Memories." Sacktor's paper with the ridiculously low sample sizes such as only four mice or six mice is passed off as a discovery, with no mention at all of the absurdly low study group sizes used. Another neuroscientist guilty of the same type of very poor research practices (such as the use of way-too-small study group sizes and utterly unreliable "freezing behavior" judgments to try to measure animal recall) is quoted as praising Sacktor's research. Again and again the article uses the word "discovery" or "discovered" for something that was not actually discovered, and again and again the article uses the word "showed" for something that was not actually showed.
Post-post-postscript: The 2025 paper here states that "the synaptic turnover rate is as high as 1% per day in the visual cortex." That is a rate of about 100% replacement every four months. If that is the typical rate at which synapses are replaced, synapses cannot be the storage place of memories lasting decades. Quoting an even higher rate of synaptic turnover, the paper here states, "A recent imaging study revealed that the synaptic turnover rate in hippocampal CA1 cells is very high, with an estimated lifetime of 1–2 weeks (Attardo et al., 2015)."

